

Cells are strongly influenced by their surroundings. Their behavior depends not only on which genes are active, but also on neighboring cells and the specific tissue environment they occupy.
Single-cell RNA sequencing, or scRNA-seq, can measure gene activity in individual cells, but the process usually removes cells from their original tissue, causing spatial information to be lost. Spatial proteomics preserves tissue location, but typically measures a much smaller set of molecular markers.
Researchers at Columbia University have developed a computational framework called ARCADIA to combine these two types of data.
Overview of the ARCADIA framework
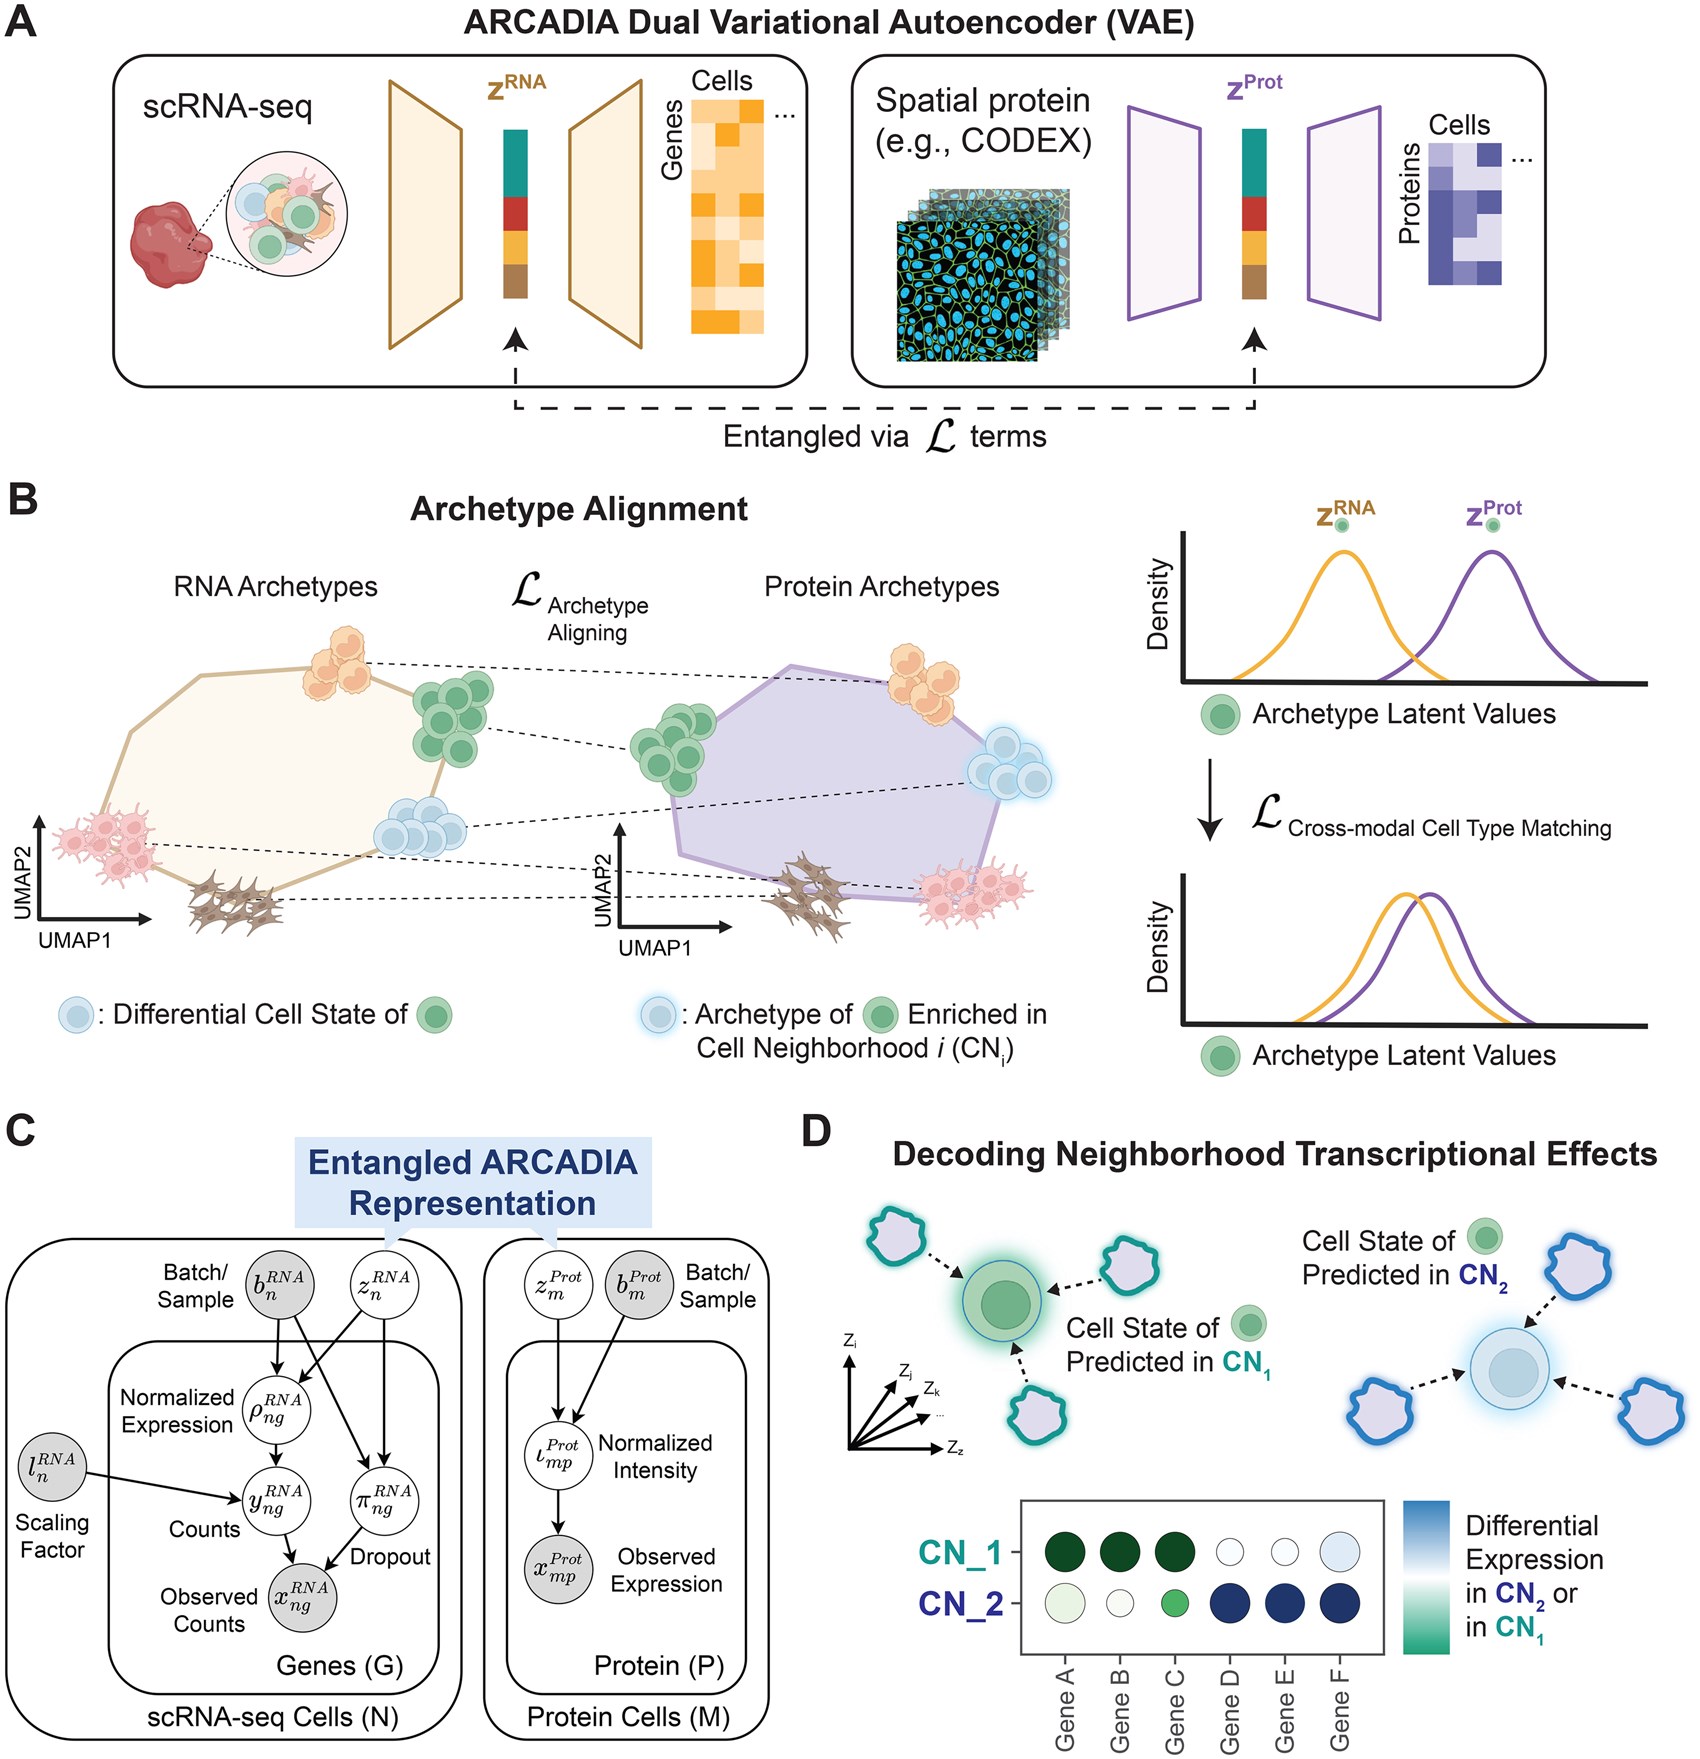
(A) Schematic of dual variational autoencoder (VAE) architecture including inputs and outputs. (B) ARCADIA aligns archetypes representing cell types/states in both modalities. Archetypes are convex combinations of extreme points on polytopes based on RNA expression (scRNA-seq) or self and neighborhood protein expression (spatial protein). Within matched archetypes, latent distributions are constrained using cross-modal matching loss to yield an entangled representation preserving cell type geometry and concordance across modalities. (C) Dual VAE graphical model. Latent variables are circles, and observed variables are shaded circles. (D) Data integration and trained dual VAEs allow for investigations into spatial dependencies of cell phenotypes including the effect of CNs on expression. CN, cell neighborhood.
ARCADIA, short for ARchetype-based Clustering and Alignment with Dual Integrative Autoencoders, does not require individual cells to be matched across datasets. Instead, it identifies representative cellular states, called archetypes, in each dataset and aligns them based on their biological composition.
The system then uses machine learning to create a shared representation of the RNA sequencing and spatial proteomics data while preserving information about nearby cells and tissue structure.
The researchers tested ARCADIA on semi-synthetic CITE-seq data and found that it performed better than existing methods designed for datasets with limited direct correspondence.
They then applied ARCADIA to independently generated scRNA-seq and CODEX spatial proteomics data from human tonsil tissue. The method reconstructed known features of tonsil organization and identified gene-expression programs associated with specific tissue environments.
In particular, ARCADIA linked B-cell maturation and T-cell activation or exhaustion with distinct spatial niches. These findings show how cells of the same general type can behave differently depending on where they are located and which cells surround them.
By combining the detailed gene-expression information from single-cell RNA sequencing with the spatial context provided by proteomics, ARCADIA could help researchers better understand how tissue environments shape cell behavior in areas such as immunology, cancer, and developmental biology.
Availability – Source code is accessible at https://github.com/azizilab/ARCADIA_public.
Rozenman B, Hoffer-Hawlik K, Djedjos N, Azizi E. (2026) ARCADIA reveals spatially dependent transcriptional programs through integration of scRNA-seq and spatial proteomics. Bioinformatics 42(Supplement_2): btag454. [article]
Related Posts




Cells are strongly influenced by their surroundings. Their behavior depends not only on which genes are active, but also on neighboring cells and the specific tissue environment they occupy.
Single-cell RNA sequencing, or scRNA-seq, can measure gene activity in individual cells, but the process usually removes cells from their original tissue, causing spatial information to be lost. Spatial proteomics preserves tissue location, but typically measures a much smaller set of molecular markers.
Researchers at Columbia University have developed a computational framework called ARCADIA to combine these two types of data.
Overview of the ARCADIA framework
(A) Schematic of dual variational autoencoder (VAE) architecture including inputs and outputs. (B) ARCADIA aligns archetypes representing cell types/states in both modalities. Archetypes are convex combinations of extreme points on polytopes based on RNA expression (scRNA-seq) or self and neighborhood protein expression (spatial protein). Within matched archetypes, latent distributions are constrained using cross-modal matching loss to yield an entangled representation preserving cell type geometry and concordance across modalities. (C) Dual VAE graphical model. Latent variables are circles, and observed variables are shaded circles. (D) Data integration and trained dual VAEs allow for investigations into spatial dependencies of cell phenotypes including the effect of CNs on expression. CN, cell neighborhood.
ARCADIA, short for ARchetype-based Clustering and Alignment with Dual Integrative Autoencoders, does not require individual cells to be matched across datasets. Instead, it identifies representative cellular states, called archetypes, in each dataset and aligns them based on their biological composition.
The system then uses machine learning to create a shared representation of the RNA sequencing and spatial proteomics data while preserving information about nearby cells and tissue structure.
The researchers tested ARCADIA on semi-synthetic CITE-seq data and found that it performed better than existing methods designed for datasets with limited direct correspondence.
They then applied ARCADIA to independently generated scRNA-seq and CODEX spatial proteomics data from human tonsil tissue. The method reconstructed known features of tonsil organization and identified gene-expression programs associated with specific tissue environments.
In particular, ARCADIA linked B-cell maturation and T-cell activation or exhaustion with distinct spatial niches. These findings show how cells of the same general type can behave differently depending on where they are located and which cells surround them.
By combining the detailed gene-expression information from single-cell RNA sequencing with the spatial context provided by proteomics, ARCADIA could help researchers better understand how tissue environments shape cell behavior in areas such as immunology, cancer, and developmental biology.
Availability – Source code is accessible at https://github.com/azizilab/ARCADIA_public.
Rozenman B, Hoffer-Hawlik K, Djedjos N, Azizi E. (2026) ARCADIA reveals spatially dependent transcriptional programs through integration of scRNA-seq and spatial proteomics. Bioinformatics 42(Supplement_2): btag454. [article]
Related Posts
Recent Posts
 Benchmarking RNA sequencing for more accurate alternative splicing analysis
 RNA Sequencing identifies new tick-borne virus that causes flu-like illness
 Eclipsebio expands its RNA characterization capabilities with multi-attribute nanopore-based sequencing assay eSTRAND RNA QC™
 Single-cell RNA sequencing links genetic mutations to cell behavior in cancer
 Small RNA sequencing reveals regulatory roles for sdRNAs in acute myeloid leukemia
 Creative Biolabs expands advanced single-cell multi-omics and RNA sequencing solutions to support biomedical discovery
RNA-Seq Products & Services


Stay Connected