By Lior Pachter –
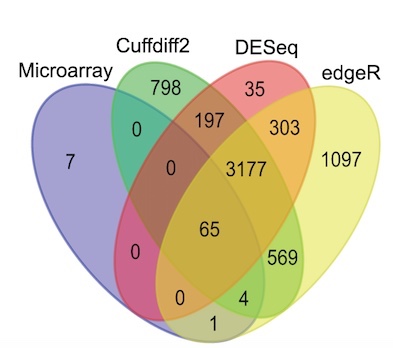
One of the maxims of computational biology is that “no two programs ever give the same result.” This is perhaps not so surprising; after all, most journals seek papers that report a significant improvement to an existing method. As a result, when developing new methods, computational biologists ensure that the results of their tools are different, specifically better (by some metric), than those of previous methods. The maxim certainly holds for RNA-Seq tools. For example, the large symmetric differences displayed in the Venn diagram below (from Zhang et al. 2014) are typical for differential expression tool benchmarks:
In a comparison of RNA-Seq quantification methods, Hayer et al. 2015 showed that methods differ even at the level of summary statistics (in Figure 7 from the paper, shown below, Pearson correlation was calculated using ground truth from a simulation):

These sort of of results are the norm in computational genomics. Finding a pair of software programs that produce identical results is about as likely as finding someone who has won the lottery… twice…. in one week. Well, it turns out there has been such a person, and here I describe the computational genomics analog of that unlikely event.
(read more…)


By Lior Pachter –
One of the maxims of computational biology is that “no two programs ever give the same result.” This is perhaps not so surprising; after all, most journals seek papers that report a significant improvement to an existing method. As a result, when developing new methods, computational biologists ensure that the results of their tools are different, specifically better (by some metric), than those of previous methods. The maxim certainly holds for RNA-Seq tools. For example, the large symmetric differences displayed in the Venn diagram below (from Zhang et al. 2014) are typical for differential expression tool benchmarks:
In a comparison of RNA-Seq quantification methods, Hayer et al. 2015 showed that methods differ even at the level of summary statistics (in Figure 7 from the paper, shown below, Pearson correlation was calculated using ground truth from a simulation):
These sort of of results are the norm in computational genomics. Finding a pair of software programs that produce identical results is about as likely as finding someone who has won the lottery… twice…. in one week. Well, it turns out there has been such a person, and here I describe the computational genomics analog of that unlikely event.
(read more…)
Related Posts
Worm’s radical transformation shows metamorphosis can change the functions of cells
RNA sequencing reveals functional chimeric mRNAs in mammalian immunity
Atlas of the brain’s striatum could guide researchers to new drug treatments
Immune cells offer insights on billion-dollar virus
A functionally integrated cross-tissue alternative splicing program during short-term calorie restriction
Dietary oxidized plant sterol shifts macrophage state to fuel aortic inflammation
Unlocking the past – new method helps gain insights into old tissue
Novel AI model trained on RNA-Seq data accurately detects key gene mutations and predicts biomarkers across 32 cancer types
Transcriptomic aging clock reveals age-related molecular patterns in opioid dependence
RNA sequencing helps predict stem cell transplant benefit in pediatric AML
Protein ‘switch’ determines whether liposarcoma cells will become aggressive
Precursor tRNAs sense temperature changes: heat stress-induced capped pre-tRNAs suppress protein synthesis
Ketamine increases neuroplasticity in female mice but not in males
Somatic mutations linked to vascular damage in progeria
Scientists map dormant cancer cells’ hideouts, opening new targets for treatment
Soluble signals released by neighboring cells direct how the human kidney is built
Genetics influence how cancer arises – and how it evolves
RNA-based testing uncovers extraordinary diversity in mutations driving lung cancer
Study offers new insights into why ex-smokers remain at elevated risk of lung disease
Learning the grammar of gene regulation
By Lior Pachter –
One of the maxims of computational biology is that “no two programs ever give the same result.” This is perhaps not so surprising; after all, most journals seek papers that report a significant improvement to an existing method. As a result, when developing new methods, computational biologists ensure that the results of their tools are different, specifically better (by some metric), than those of previous methods. The maxim certainly holds for RNA-Seq tools. For example, the large symmetric differences displayed in the Venn diagram below (from Zhang et al. 2014) are typical for differential expression tool benchmarks:
In a comparison of RNA-Seq quantification methods, Hayer et al. 2015 showed that methods differ even at the level of summary statistics (in Figure 7 from the paper, shown below, Pearson correlation was calculated using ground truth from a simulation):
These sort of of results are the norm in computational genomics. Finding a pair of software programs that produce identical results is about as likely as finding someone who has won the lottery… twice…. in one week. Well, it turns out there has been such a person, and here I describe the computational genomics analog of that unlikely event.
(read more…)
Related Posts
Worm’s radical transformation shows metamorphosis can change the functions of cells
RNA sequencing reveals functional chimeric mRNAs in mammalian immunity
Atlas of the brain’s striatum could guide researchers to new drug treatments
Immune cells offer insights on billion-dollar virus
A functionally integrated cross-tissue alternative splicing program during short-term calorie restriction
Dietary oxidized plant sterol shifts macrophage state to fuel aortic inflammation
Unlocking the past – new method helps gain insights into old tissue
Novel AI model trained on RNA-Seq data accurately detects key gene mutations and predicts biomarkers across 32 cancer types
Transcriptomic aging clock reveals age-related molecular patterns in opioid dependence
RNA sequencing helps predict stem cell transplant benefit in pediatric AML
Protein ‘switch’ determines whether liposarcoma cells will become aggressive
Precursor tRNAs sense temperature changes: heat stress-induced capped pre-tRNAs suppress protein synthesis
Ketamine increases neuroplasticity in female mice but not in males
Somatic mutations linked to vascular damage in progeria
Scientists map dormant cancer cells’ hideouts, opening new targets for treatment
Soluble signals released by neighboring cells direct how the human kidney is built
Genetics influence how cancer arises – and how it evolves
RNA-based testing uncovers extraordinary diversity in mutations driving lung cancer
Study offers new insights into why ex-smokers remain at elevated risk of lung disease
Learning the grammar of gene regulation
Stay Connected
Submit a Post to the Blog
Recent Posts
Subscribe to the RNA-Seq Blog
RNA-Seq Products & Services