

Researchers at the University of Southampton have developed a cutting-edge tool that can rapidly identify and characterize tumor-related immunoglobulin genes directly from RNA sequencing (RNA-seq) data. This groundbreaking technique has the potential to significantly enhance our understanding of B cell-related cancers like chronic lymphocytic leukemia (CLL) and lymphomas, ultimately improving diagnosis, prognosis, and treatment options.
Immunoglobulin Genes and Their Role in B Cell Tumors
B cells are a type of white blood cell that plays a crucial role in the immune system by producing antibodies to fight infections. Each B cell has a unique immunoglobulin (Ig) gene that encodes the antibodies it produces. These Ig genes are crucial to understanding the behavior of B cell tumors. In cancers of B cells, like CLL and lymphomas, analyzing these Ig genes can provide valuable insights into the tumor’s origin, the type of mutation it carries, and how aggressive it might be.
Traditional methods like PCR and Sanger sequencing, although effective, are slow and often unable to capture the full complexity of the immunoglobulin gene sequence. These methods also rely on identifying a dominant sequence or a small number of subclonal sequences, which limits their ability to analyze the complete landscape of B cell tumors.
The Power of RNA Sequencing in Cancer Research
With the advent of RNA sequencing, scientists can now analyze the entire transcriptome of a tumor, including the immunoglobulin genes, in a much faster and more comprehensive way. This approach allows researchers to capture the full immunoglobulin gene sequence—both templated and non-templated regions—from RNA samples extracted directly from tumors. However, this new wealth of data requires advanced tools to manage and interpret.
IgSeqR is a user-friendly, high-performance protocol that enables the rapid (~1 hour) de novo assembly and accurate characterization of the full immunoglobulin transcript sequence from RNA-seq data. This includes both the leader region and the constant region of the immunoglobulin gene, which is crucial for understanding the tumor’s molecular features.
Schematic representation of the IgSeqR pipeline
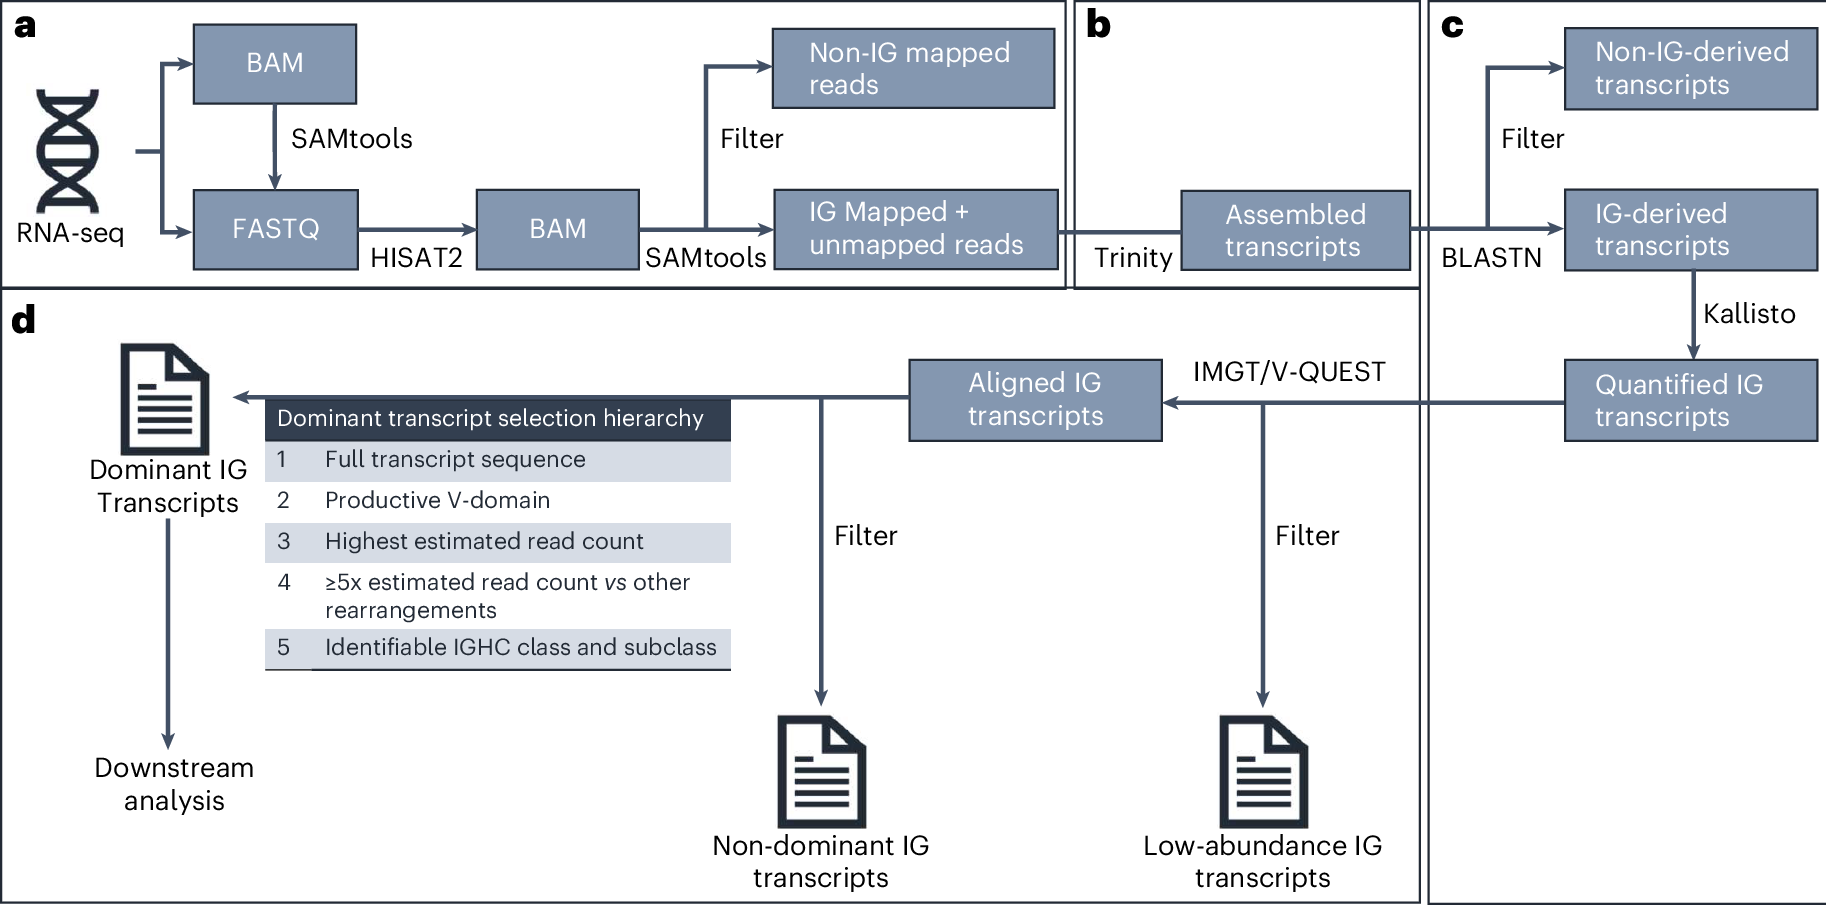
The procedure for running IgSeqR is divided into four key stages. The first stage (a) is data pre-processing, in which RNA-seq data can be supplied in either BAM or FASTQ format. If starting from a BAM file, reads are first extracted in FASTQ format by using the Samtools fastq command. The data are re-aligned to a reference transcriptome by HISAT2, producing a BAM file that is filtered to retain reads mapping to immunoglobulin gene coordinates and reads unable to be mapped to the reference. The second stage (b) is de novo transcriptome assembly, in which Trinity is used to assemble transcripts de novo from the filtered BAM file. The third stage (c) is immunoglobulin transcript selection and quantification, in which the assembled transcripts are run through a BLAST query to identify transcripts overlapping immunoglobulin reference sequences. The abundance of the immunoglobulin-derived transcripts is then estimated by using Kallisto pseudoalignment. The fourth stage (d) is immunoglobulin transcript annotation and interpretation, in which the five most-abundant transcripts by transcript per million are then run through IMGT/V-QUEST for immunoglobulin alignment and annotation, which is used to recover the dominant immunoglobulin transcript originating from the putative tumor immunoglobulin gene by using a five-step hierarchical selection process.
Beyond Just Gene Identification: Understanding Tumor Glycosylation
What sets IgSeqR apart is its ability to provide not just sequence data, but also the ability to examine the amino acid sequences of the immunoglobulin genes for important post-translational modifications like glycosylation. Glycosylation refers to the addition of sugar molecules to proteins, and these sugar modifications can affect how the protein functions. In some lymphomas, specific glycosylation patterns are associated with tumor progression and immune evasion, making them valuable biomarkers for prognosis.
With IgSeqR, scientists can now link the immunoglobulin sequence data to the overall tumor transcriptome, allowing for deeper insights into the tumor’s molecular environment. This integrated approach can help refine diagnoses and predict how the tumor might respond to treatments. It also opens the door for personalized medicine strategies, where therapies could be tailored to the specific genetic and molecular makeup of an individual’s tumor.
Improving Diagnosis and Prognosis for B Cell Tumors
The IgSeqR tool is a major step forward in cancer genomics because it allows for a more comprehensive understanding of the tumor at the molecular level. By combining immunoglobulin gene data with other tumor characteristics, researchers can gain insights into tumor origin, progression, and response to treatment. This can be especially helpful in B cell cancers, which can be highly heterogeneous and challenging to treat.
For example, IgSeqR can be used to analyze specific features of tumors that are associated with treatment resistance, such as mutations in immunoglobulin genes or altered glycosylation patterns. This information could help doctors make more informed decisions about treatment options, potentially leading to better outcomes for patients.
A Tool for the Future of Cancer Research and Treatment
As more B cell tumors are analyzed using IgSeqR, it’s likely that the tool will play a crucial role in advancing cancer research. Its ability to provide accurate, high-quality immunoglobulin gene sequences directly from RNA sequencing data could revolutionize the way we approach B cell tumors, from early detection and diagnosis to targeted therapy and personalized treatment plans.
Availability – The code for IgSeqR is available on GitHub at https://github.com/ForconiLab/IgSeqR/releases/tag/v1.0.1.
Bryant D, Sale B, Chiodin G, et al. (2025) Identification, assembly and characterization of tumor immunoglobulin transcripts from RNA sequencing data using IgSeqR. Nat Protoc [Epub ahead of print]. [article]
Related Posts



Researchers at the University of Southampton have developed a cutting-edge tool that can rapidly identify and characterize tumor-related immunoglobulin genes directly from RNA sequencing (RNA-seq) data. This groundbreaking technique has the potential to significantly enhance our understanding of B cell-related cancers like chronic lymphocytic leukemia (CLL) and lymphomas, ultimately improving diagnosis, prognosis, and treatment options.
Immunoglobulin Genes and Their Role in B Cell Tumors
B cells are a type of white blood cell that plays a crucial role in the immune system by producing antibodies to fight infections. Each B cell has a unique immunoglobulin (Ig) gene that encodes the antibodies it produces. These Ig genes are crucial to understanding the behavior of B cell tumors. In cancers of B cells, like CLL and lymphomas, analyzing these Ig genes can provide valuable insights into the tumor’s origin, the type of mutation it carries, and how aggressive it might be.
Traditional methods like PCR and Sanger sequencing, although effective, are slow and often unable to capture the full complexity of the immunoglobulin gene sequence. These methods also rely on identifying a dominant sequence or a small number of subclonal sequences, which limits their ability to analyze the complete landscape of B cell tumors.
The Power of RNA Sequencing in Cancer Research
With the advent of RNA sequencing, scientists can now analyze the entire transcriptome of a tumor, including the immunoglobulin genes, in a much faster and more comprehensive way. This approach allows researchers to capture the full immunoglobulin gene sequence—both templated and non-templated regions—from RNA samples extracted directly from tumors. However, this new wealth of data requires advanced tools to manage and interpret.
IgSeqR is a user-friendly, high-performance protocol that enables the rapid (~1 hour) de novo assembly and accurate characterization of the full immunoglobulin transcript sequence from RNA-seq data. This includes both the leader region and the constant region of the immunoglobulin gene, which is crucial for understanding the tumor’s molecular features.
Schematic representation of the IgSeqR pipeline
The procedure for running IgSeqR is divided into four key stages. The first stage (a) is data pre-processing, in which RNA-seq data can be supplied in either BAM or FASTQ format. If starting from a BAM file, reads are first extracted in FASTQ format by using the Samtools fastq command. The data are re-aligned to a reference transcriptome by HISAT2, producing a BAM file that is filtered to retain reads mapping to immunoglobulin gene coordinates and reads unable to be mapped to the reference. The second stage (b) is de novo transcriptome assembly, in which Trinity is used to assemble transcripts de novo from the filtered BAM file. The third stage (c) is immunoglobulin transcript selection and quantification, in which the assembled transcripts are run through a BLAST query to identify transcripts overlapping immunoglobulin reference sequences. The abundance of the immunoglobulin-derived transcripts is then estimated by using Kallisto pseudoalignment. The fourth stage (d) is immunoglobulin transcript annotation and interpretation, in which the five most-abundant transcripts by transcript per million are then run through IMGT/V-QUEST for immunoglobulin alignment and annotation, which is used to recover the dominant immunoglobulin transcript originating from the putative tumor immunoglobulin gene by using a five-step hierarchical selection process.
Beyond Just Gene Identification: Understanding Tumor Glycosylation
What sets IgSeqR apart is its ability to provide not just sequence data, but also the ability to examine the amino acid sequences of the immunoglobulin genes for important post-translational modifications like glycosylation. Glycosylation refers to the addition of sugar molecules to proteins, and these sugar modifications can affect how the protein functions. In some lymphomas, specific glycosylation patterns are associated with tumor progression and immune evasion, making them valuable biomarkers for prognosis.
With IgSeqR, scientists can now link the immunoglobulin sequence data to the overall tumor transcriptome, allowing for deeper insights into the tumor’s molecular environment. This integrated approach can help refine diagnoses and predict how the tumor might respond to treatments. It also opens the door for personalized medicine strategies, where therapies could be tailored to the specific genetic and molecular makeup of an individual’s tumor.
Improving Diagnosis and Prognosis for B Cell Tumors
The IgSeqR tool is a major step forward in cancer genomics because it allows for a more comprehensive understanding of the tumor at the molecular level. By combining immunoglobulin gene data with other tumor characteristics, researchers can gain insights into tumor origin, progression, and response to treatment. This can be especially helpful in B cell cancers, which can be highly heterogeneous and challenging to treat.
For example, IgSeqR can be used to analyze specific features of tumors that are associated with treatment resistance, such as mutations in immunoglobulin genes or altered glycosylation patterns. This information could help doctors make more informed decisions about treatment options, potentially leading to better outcomes for patients.
A Tool for the Future of Cancer Research and Treatment
As more B cell tumors are analyzed using IgSeqR, it’s likely that the tool will play a crucial role in advancing cancer research. Its ability to provide accurate, high-quality immunoglobulin gene sequences directly from RNA sequencing data could revolutionize the way we approach B cell tumors, from early detection and diagnosis to targeted therapy and personalized treatment plans.
Availability – The code for IgSeqR is available on GitHub at https://github.com/ForconiLab/IgSeqR/releases/tag/v1.0.1.
Bryant D, Sale B, Chiodin G, et al. (2025) Identification, assembly and characterization of tumor immunoglobulin transcripts from RNA sequencing data using IgSeqR. Nat Protoc [Epub ahead of print]. [article]
Related Posts
Recent Posts
 Benchmarking RNA sequencing for more accurate alternative splicing analysis
 RNA Sequencing identifies new tick-borne virus that causes flu-like illness
 Eclipsebio expands its RNA characterization capabilities with multi-attribute nanopore-based sequencing assay eSTRAND RNA QC™
 Single-cell RNA sequencing links genetic mutations to cell behavior in cancer
 Small RNA sequencing reveals regulatory roles for sdRNAs in acute myeloid leukemia
 Creative Biolabs expands advanced single-cell multi-omics and RNA sequencing solutions to support biomedical discovery
RNA-Seq Products & Services


Stay Connected