

Understanding how genetic differences influence disease and aging at the cellular level is one of the biggest challenges in modern biology. While large genetic studies called genome-wide association studies, or GWASs, can point to DNA variants linked to conditions like Parkinson’s disease and immune disorders, they do not explain how these variants exert their effects in specific cell types. A team led by researchers at the University of Pennsylvania have developed a powerful new method, called scMORE, that brings together several types of data to solve this puzzle.
scMORE combines single-cell RNA sequencing and chromatin accessibility information with GWAS summary data. Single-cell RNA sequencing tells us which genes are active in each cell, while chromatin accessibility data reveals which parts of the genome are open for regulation. By integrating these datasets with genetic risk signals from GWASs, scMORE identifies eRegulons, groups of genes controlled by shared transcription factors that are linked to genetic variants associated with disease or aging traits.
Overview of the scMORE
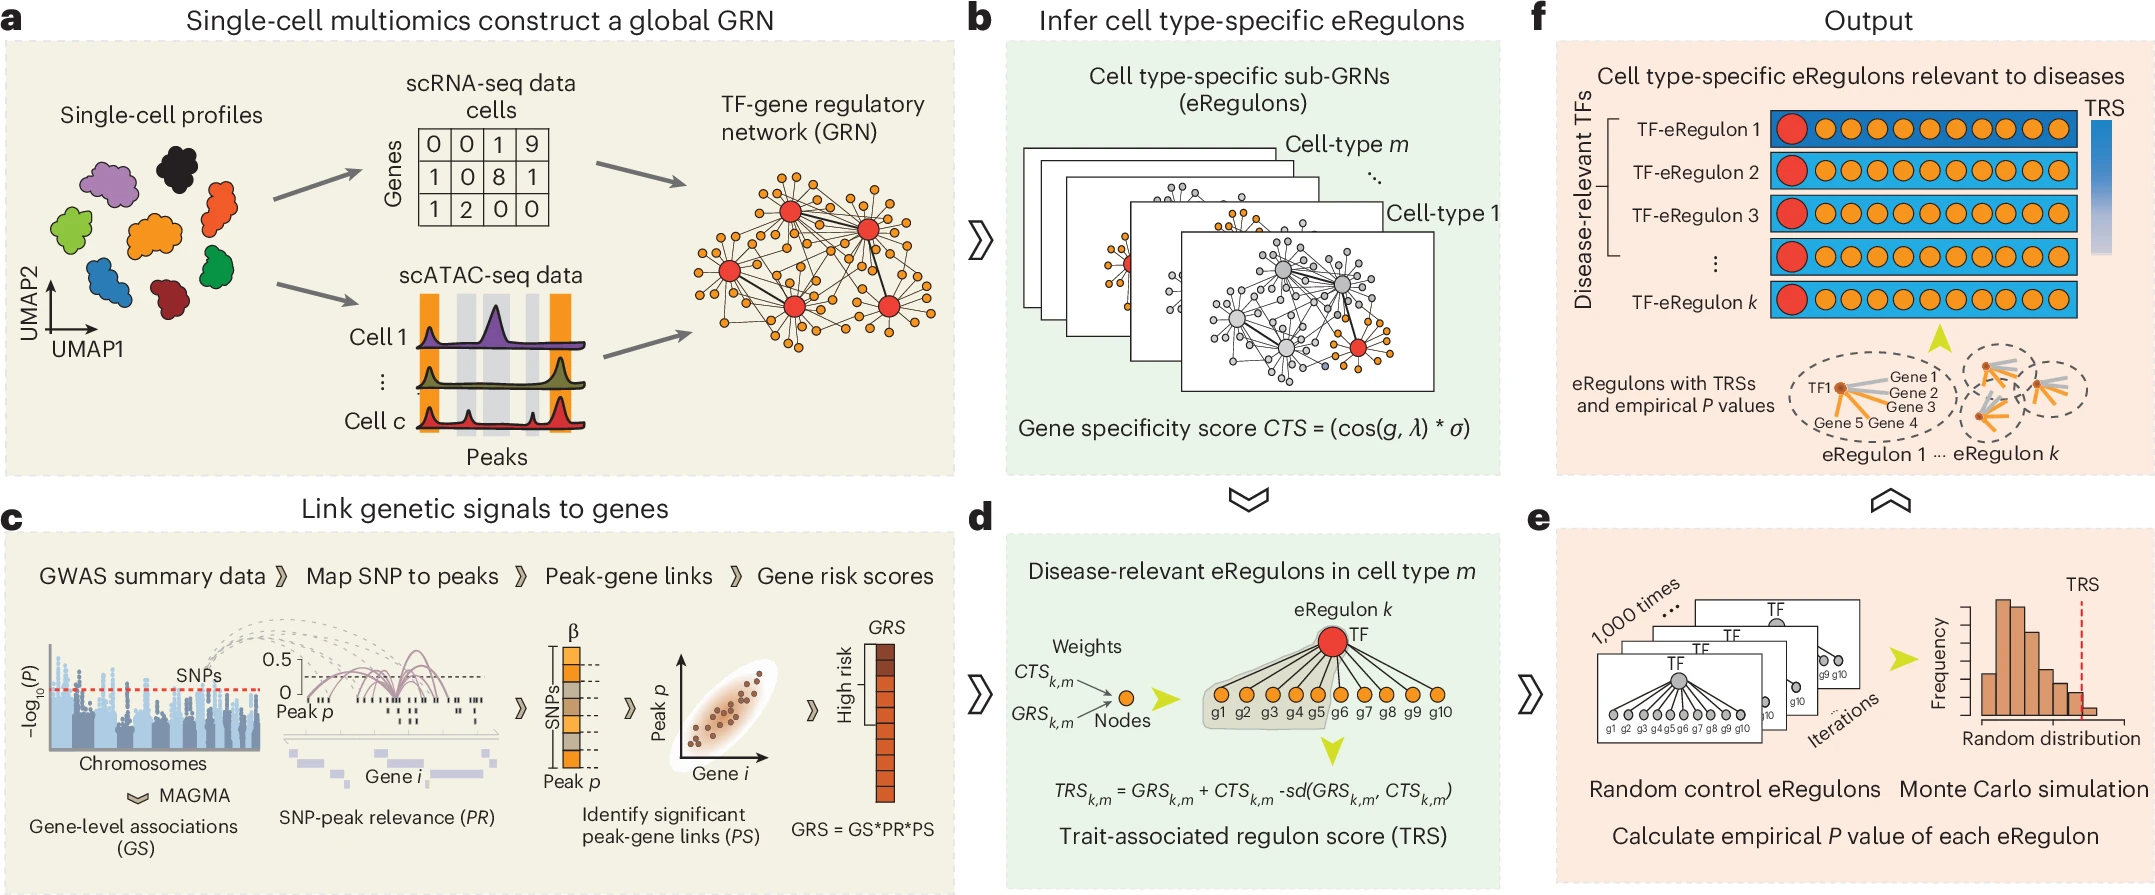
a, Utilizing single-cell multimodal measurements to construct a global TF-GRN based on a GLM. b, A modified cosine similarity method is applied to infer the specificity score of each TF and its target genes (CTS) within a sub-GRN (eRegulon) in a specific cell type. c, Genetic association signals from GWAS summary statistics are mapped to peaks (PR), linked to corresponding genes (PS) and disease-specific genes (GS) are identified using MAGMA or FUMA gene-scoring methods. The GRS for each node is calculated as the product of GS, PR and PS, representing the genetic correlation of the node with the trait. d, The TRS is calculated for each eRegulon in a particular cell type by integrating the GRS and CTS scores. The s.d. term is included as a penalty to control the deviation between GWAS signals and gene expression. e, MC Model: 1,000 sets of matched control eRegulons are generated for each eRegulon in a given cell type. MC sampling is then used to calculate a P value for each eRegulon based on the empirical distribution. f, scMORE outputs. The outputs include (1) TRS, GRS and CTS values for each eRegulon associated with a specific disease in a given cell type; (2) MC-based empirical P values for each eRegulon within the specific cellular context.
The strength of scMORE lies in its ability to pinpoint which regulatory networks are active in specific cell types and how genetic variants influence those networks. The researchers tested scMORE using data from 31 human traits related to immune function and aging, including Parkinson’s disease. In the human midbrain, scMORE uncovered 77 eRegulons connected to aging and Parkinson’s disease across seven distinct brain cell types. Remarkably, the analysis also revealed differences between males and females in how these regulatory networks behave in neurons affected by Parkinson’s disease compared with younger and older individuals without the disease.
By linking DNA variants to cell-type-specific regulatory activity, scMORE provides a clearer picture of how genetic risks shape biological networks in health and disease. This framework helps researchers move beyond simply knowing that a variant is associated with a condition to understanding how it influences gene regulation in particular cells. Such insights can guide the development of new therapeutic strategies and improve our understanding of complex diseases rooted in aging and immune processes.
Availability – The scMORE R package, an open-source software, has been developed and is freely accessible on the GitHub repository (https://github.com/mayunlong89/scMORE).
Ma Y, Yao Y, Zhou Y, Dai W, Li J, Gui Y, Sun H, Zhu Z, Jiang D, Chen C, Deng C, Huang Y, Han H, Zhou J, Su J. (2025) Integrating polygenic signals and single-cell multiomics identifies cell-type-specific regulomes critical for immune- and aging-related diseases. Nature Aging [Epub ahead of print]. [article]
Related Posts





Understanding how genetic differences influence disease and aging at the cellular level is one of the biggest challenges in modern biology. While large genetic studies called genome-wide association studies, or GWASs, can point to DNA variants linked to conditions like Parkinson’s disease and immune disorders, they do not explain how these variants exert their effects in specific cell types. A team led by researchers at the University of Pennsylvania have developed a powerful new method, called scMORE, that brings together several types of data to solve this puzzle.
scMORE combines single-cell RNA sequencing and chromatin accessibility information with GWAS summary data. Single-cell RNA sequencing tells us which genes are active in each cell, while chromatin accessibility data reveals which parts of the genome are open for regulation. By integrating these datasets with genetic risk signals from GWASs, scMORE identifies eRegulons, groups of genes controlled by shared transcription factors that are linked to genetic variants associated with disease or aging traits.
Overview of the scMORE
a, Utilizing single-cell multimodal measurements to construct a global TF-GRN based on a GLM. b, A modified cosine similarity method is applied to infer the specificity score of each TF and its target genes (CTS) within a sub-GRN (eRegulon) in a specific cell type. c, Genetic association signals from GWAS summary statistics are mapped to peaks (PR), linked to corresponding genes (PS) and disease-specific genes (GS) are identified using MAGMA or FUMA gene-scoring methods. The GRS for each node is calculated as the product of GS, PR and PS, representing the genetic correlation of the node with the trait. d, The TRS is calculated for each eRegulon in a particular cell type by integrating the GRS and CTS scores. The s.d. term is included as a penalty to control the deviation between GWAS signals and gene expression. e, MC Model: 1,000 sets of matched control eRegulons are generated for each eRegulon in a given cell type. MC sampling is then used to calculate a P value for each eRegulon based on the empirical distribution. f, scMORE outputs. The outputs include (1) TRS, GRS and CTS values for each eRegulon associated with a specific disease in a given cell type; (2) MC-based empirical P values for each eRegulon within the specific cellular context.
The strength of scMORE lies in its ability to pinpoint which regulatory networks are active in specific cell types and how genetic variants influence those networks. The researchers tested scMORE using data from 31 human traits related to immune function and aging, including Parkinson’s disease. In the human midbrain, scMORE uncovered 77 eRegulons connected to aging and Parkinson’s disease across seven distinct brain cell types. Remarkably, the analysis also revealed differences between males and females in how these regulatory networks behave in neurons affected by Parkinson’s disease compared with younger and older individuals without the disease.
By linking DNA variants to cell-type-specific regulatory activity, scMORE provides a clearer picture of how genetic risks shape biological networks in health and disease. This framework helps researchers move beyond simply knowing that a variant is associated with a condition to understanding how it influences gene regulation in particular cells. Such insights can guide the development of new therapeutic strategies and improve our understanding of complex diseases rooted in aging and immune processes.
Availability – The scMORE R package, an open-source software, has been developed and is freely accessible on the GitHub repository (https://github.com/mayunlong89/scMORE).
Ma Y, Yao Y, Zhou Y, Dai W, Li J, Gui Y, Sun H, Zhu Z, Jiang D, Chen C, Deng C, Huang Y, Han H, Zhou J, Su J. (2025) Integrating polygenic signals and single-cell multiomics identifies cell-type-specific regulomes critical for immune- and aging-related diseases. Nature Aging [Epub ahead of print]. [article]
Related Posts
Recent Posts
 Benchmarking RNA sequencing for more accurate alternative splicing analysis
 RNA Sequencing identifies new tick-borne virus that causes flu-like illness
 Eclipsebio expands its RNA characterization capabilities with multi-attribute nanopore-based sequencing assay eSTRAND RNA QC™
 Single-cell RNA sequencing links genetic mutations to cell behavior in cancer
 Small RNA sequencing reveals regulatory roles for sdRNAs in acute myeloid leukemia
 Creative Biolabs expands advanced single-cell multi-omics and RNA sequencing solutions to support biomedical discovery
RNA-Seq Products & Services


Stay Connected