A groundbreaking study led by researchers at Beth Israel Deaconess Medical Center (BIDMC) and Boston Children’s Hospital reveals how tumors grow by avoiding the immune system. The study reveals how genes in tumors edit themselves to escape the immune system’s detection. The research identifies, for the first time, the actual genes that are silenced by tumors, offering a roadmap for better immunotherapies.
The researchers studied breast cancer in a mouse model, using genome-wide single-cell RNA sequencing to identify which genes were edited by tumors as they developed. The team found that tumors use epigenetic modification, particularly DNA methylation, to suppress genes involved in the innate immune response—the body’s first line of defense against pathogens and disease. This silencing is how most tumors evade current immunotherapies, such as CAR-T cells and checkpoint inhibitors. However, the scientists also found that the FDA-approved drug decitabine, a chemotherapy that slows cancer cell growth, can reverse these gene edits. At low doses, decitabine reactivates the immune response, leading to a reduction in tumor growth by boosting the presence of immune cells that attack the tumor.
Cell mapping by scRNA-seq of early and late tumors in a breast cancer GEMM
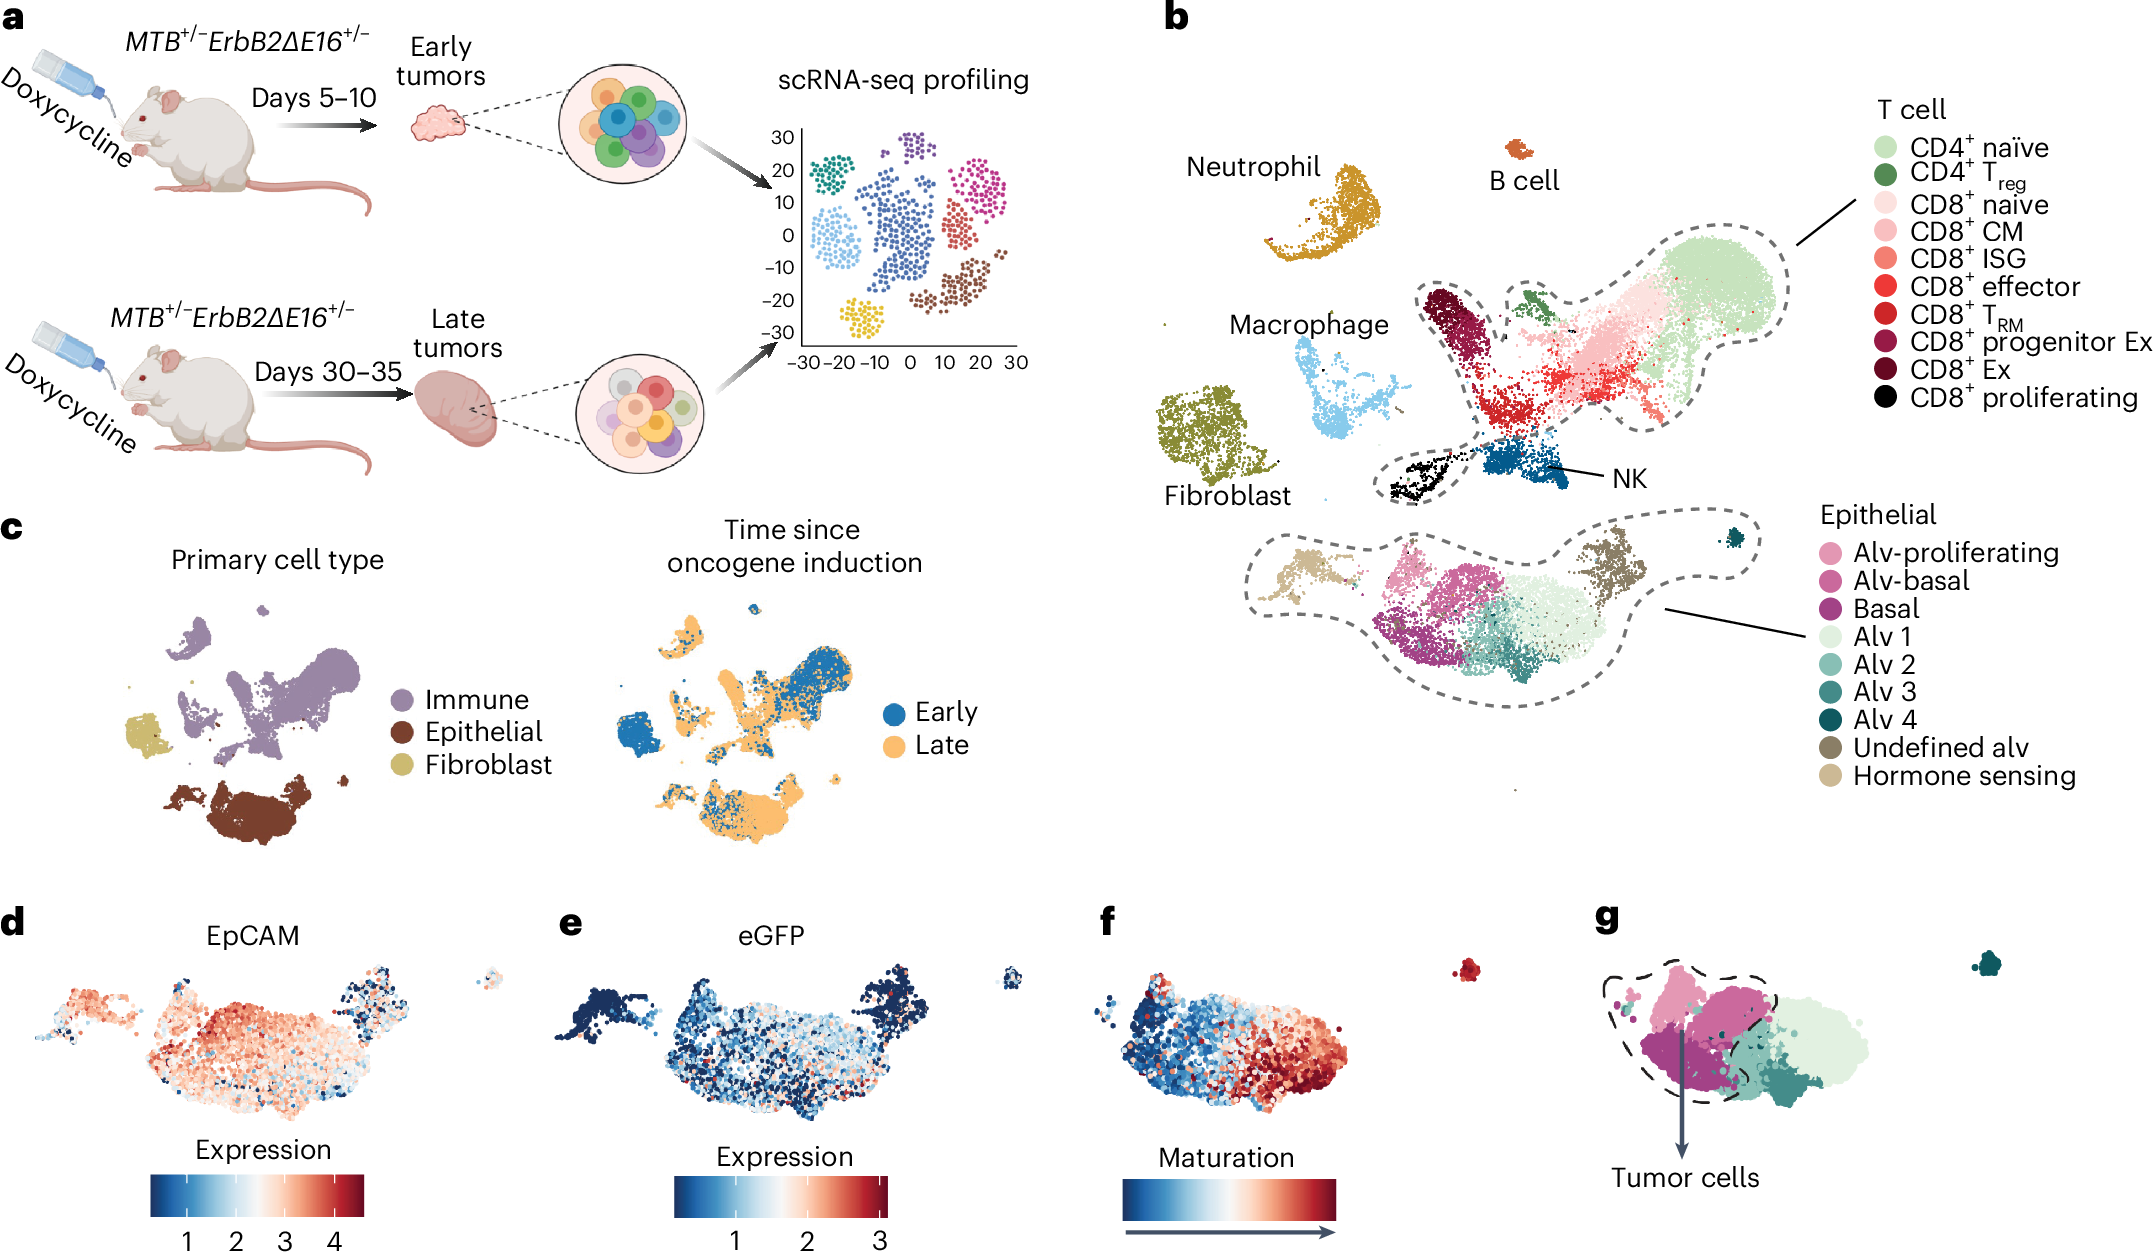
a, Experimental set up: 8-week-old female ErbB2ΔEx16 transgenic mice were given doxycycline in the drinking water to induce breast tumors, which were harvested at early (days 5–10, n = 4) or late (days 30–35, n = 4) time points and dissociated into single cells, which were analyzed for scRNA-seq. b, Uniform Manifold Approximation and Projection (UMAP) plot of 24,798 cells and 20,499 genes in early and late GEMM libraries: 23 primary cell subtypes were identified using unsupervised clustering algorithms and manual inspection of known cell markers. c, UMAP plots showing primary cell types (left) or distribution of cells in early and late tumors (right). Primary cell types were EpCAM+ epithelial cells (early tumor n = 927, late tumor n = 6,582), CD45+ immune cells (early tumor n = 7,152, late tumor n = 8,539) and FAP+/DCN+ fibroblasts (early tumor n = 1,546, late tumor n = 52). d–g, Epithelial clusters were classified as tumor and nontumor (g) based on expression of EpCAM (d) and eGFP (e) and maturation status of EpCAM+eGFP+ cells (f) as determined by CytoTRACE.
The findings suggest that gene editing might be a common defense mechanism across various cancers. Further research is needed to explore how these insights apply to other solid tumors like lung and ovarian cancer. Future studies will include studying gene editing in patients with genetic predispositions to cancer, especially at early tumor stages and during metastasis.
“Our research sheds light on the sophisticated strategies tumors use to hide from the immune system,” said Winston Hide, PhD, a lead author on the study and co-Director of the Non-Coding RNA Precision Diagnostics and Therapeutics Core Facility at BIDMC. “By identifying the specific genes that tumors silence to evade detection, we open the door to new therapeutic approaches. These findings could lead to the development of more targeted immunotherapies, potentially improving outcomes for patients with cancer.”
Source – Beth Israel Deaconess Medical Center


A groundbreaking study led by researchers at Beth Israel Deaconess Medical Center (BIDMC) and Boston Children’s Hospital reveals how tumors grow by avoiding the immune system. The study reveals how genes in tumors edit themselves to escape the immune system’s detection. The research identifies, for the first time, the actual genes that are silenced by tumors, offering a roadmap for better immunotherapies.
The researchers studied breast cancer in a mouse model, using genome-wide single-cell RNA sequencing to identify which genes were edited by tumors as they developed. The team found that tumors use epigenetic modification, particularly DNA methylation, to suppress genes involved in the innate immune response—the body’s first line of defense against pathogens and disease. This silencing is how most tumors evade current immunotherapies, such as CAR-T cells and checkpoint inhibitors. However, the scientists also found that the FDA-approved drug decitabine, a chemotherapy that slows cancer cell growth, can reverse these gene edits. At low doses, decitabine reactivates the immune response, leading to a reduction in tumor growth by boosting the presence of immune cells that attack the tumor.
Cell mapping by scRNA-seq of early and late tumors in a breast cancer GEMM
a, Experimental set up: 8-week-old female ErbB2ΔEx16 transgenic mice were given doxycycline in the drinking water to induce breast tumors, which were harvested at early (days 5–10, n = 4) or late (days 30–35, n = 4) time points and dissociated into single cells, which were analyzed for scRNA-seq. b, Uniform Manifold Approximation and Projection (UMAP) plot of 24,798 cells and 20,499 genes in early and late GEMM libraries: 23 primary cell subtypes were identified using unsupervised clustering algorithms and manual inspection of known cell markers. c, UMAP plots showing primary cell types (left) or distribution of cells in early and late tumors (right). Primary cell types were EpCAM+ epithelial cells (early tumor n = 927, late tumor n = 6,582), CD45+ immune cells (early tumor n = 7,152, late tumor n = 8,539) and FAP+/DCN+ fibroblasts (early tumor n = 1,546, late tumor n = 52). d–g, Epithelial clusters were classified as tumor and nontumor (g) based on expression of EpCAM (d) and eGFP (e) and maturation status of EpCAM+eGFP+ cells (f) as determined by CytoTRACE.
The findings suggest that gene editing might be a common defense mechanism across various cancers. Further research is needed to explore how these insights apply to other solid tumors like lung and ovarian cancer. Future studies will include studying gene editing in patients with genetic predispositions to cancer, especially at early tumor stages and during metastasis.
Source – Beth Israel Deaconess Medical Center
Zhang Y, Naderi Yeganeh P, Zhang H, Wang SY, Li Z, Gu B, Lee DJ, Zhang Z, Ploumakis A, Shi M, Wu H, Greer EL, Hide W, Lieberman J. (2024) Tumor editing suppresses innate and adaptive antitumor immunity and is reversed by inhibiting DNA methylation. Nat Immunol [Epub ahead of print]. [article]
Related Posts
Single-cell and single-embryo RNA sequencing
RNA sequencing reveals functional chimeric mRNAs in mammalian immunity
Deep learning improves microRNA target prediction from sequence
Atlas of the brain’s striatum could guide researchers to new drug treatments
scLS – a computationally efficient differentially expressed gene detection algorithm
Spatial mapping of RNA turnover kinetics in the mouse brain
Immune cells offer insights on billion-dollar virus
SPIDER improves spatial transcriptomics data using single-cell RNA sequencing
Ultrafast and reference-free sequence discovery in single-cell data
ARCADIA combines RNA sequencing and spatial proteomics to reveal how tissue location shapes cell behavior
An end-to-end computational framework for “Record-seq” transcriptional recording data
A functionally integrated cross-tissue alternative splicing program during short-term calorie restriction
ExoShorkie – predicting RNA-seq coverage of exogenous genomes in yeast by transfer learning
Dietary oxidized plant sterol shifts macrophage state to fuel aortic inflammation
Bonsai reconstructs tree representations for distortion-free visualization and exploration of high-dimensional data
MiRQuery – a user-friendly web app for the interactive analysis and visualization of microRNA sequencing data
RNA sequencing resolves cryptic pathogenic variants in mitochondrial disease
Unlocking the past – new method helps gain insights into old tissue
New RNA sequencing model improves sequencing depth planning for UMI transcriptomics
Combining RNA sequencing and pathology images identifies glioblastoma subgroups linked to survival
A groundbreaking study led by researchers at Beth Israel Deaconess Medical Center (BIDMC) and Boston Children’s Hospital reveals how tumors grow by avoiding the immune system. The study reveals how genes in tumors edit themselves to escape the immune system’s detection. The research identifies, for the first time, the actual genes that are silenced by tumors, offering a roadmap for better immunotherapies.
The researchers studied breast cancer in a mouse model, using genome-wide single-cell RNA sequencing to identify which genes were edited by tumors as they developed. The team found that tumors use epigenetic modification, particularly DNA methylation, to suppress genes involved in the innate immune response—the body’s first line of defense against pathogens and disease. This silencing is how most tumors evade current immunotherapies, such as CAR-T cells and checkpoint inhibitors. However, the scientists also found that the FDA-approved drug decitabine, a chemotherapy that slows cancer cell growth, can reverse these gene edits. At low doses, decitabine reactivates the immune response, leading to a reduction in tumor growth by boosting the presence of immune cells that attack the tumor.
Cell mapping by scRNA-seq of early and late tumors in a breast cancer GEMM
a, Experimental set up: 8-week-old female ErbB2ΔEx16 transgenic mice were given doxycycline in the drinking water to induce breast tumors, which were harvested at early (days 5–10, n = 4) or late (days 30–35, n = 4) time points and dissociated into single cells, which were analyzed for scRNA-seq. b, Uniform Manifold Approximation and Projection (UMAP) plot of 24,798 cells and 20,499 genes in early and late GEMM libraries: 23 primary cell subtypes were identified using unsupervised clustering algorithms and manual inspection of known cell markers. c, UMAP plots showing primary cell types (left) or distribution of cells in early and late tumors (right). Primary cell types were EpCAM+ epithelial cells (early tumor n = 927, late tumor n = 6,582), CD45+ immune cells (early tumor n = 7,152, late tumor n = 8,539) and FAP+/DCN+ fibroblasts (early tumor n = 1,546, late tumor n = 52). d–g, Epithelial clusters were classified as tumor and nontumor (g) based on expression of EpCAM (d) and eGFP (e) and maturation status of EpCAM+eGFP+ cells (f) as determined by CytoTRACE.
The findings suggest that gene editing might be a common defense mechanism across various cancers. Further research is needed to explore how these insights apply to other solid tumors like lung and ovarian cancer. Future studies will include studying gene editing in patients with genetic predispositions to cancer, especially at early tumor stages and during metastasis.
Source – Beth Israel Deaconess Medical Center
Zhang Y, Naderi Yeganeh P, Zhang H, Wang SY, Li Z, Gu B, Lee DJ, Zhang Z, Ploumakis A, Shi M, Wu H, Greer EL, Hide W, Lieberman J. (2024) Tumor editing suppresses innate and adaptive antitumor immunity and is reversed by inhibiting DNA methylation. Nat Immunol [Epub ahead of print]. [article]
Related Posts
Single-cell and single-embryo RNA sequencing
RNA sequencing reveals functional chimeric mRNAs in mammalian immunity
Deep learning improves microRNA target prediction from sequence
Atlas of the brain’s striatum could guide researchers to new drug treatments
scLS – a computationally efficient differentially expressed gene detection algorithm
Spatial mapping of RNA turnover kinetics in the mouse brain
Immune cells offer insights on billion-dollar virus
SPIDER improves spatial transcriptomics data using single-cell RNA sequencing
Ultrafast and reference-free sequence discovery in single-cell data
ARCADIA combines RNA sequencing and spatial proteomics to reveal how tissue location shapes cell behavior
An end-to-end computational framework for “Record-seq” transcriptional recording data
A functionally integrated cross-tissue alternative splicing program during short-term calorie restriction
ExoShorkie – predicting RNA-seq coverage of exogenous genomes in yeast by transfer learning
Dietary oxidized plant sterol shifts macrophage state to fuel aortic inflammation
Bonsai reconstructs tree representations for distortion-free visualization and exploration of high-dimensional data
MiRQuery – a user-friendly web app for the interactive analysis and visualization of microRNA sequencing data
RNA sequencing resolves cryptic pathogenic variants in mitochondrial disease
Unlocking the past – new method helps gain insights into old tissue
New RNA sequencing model improves sequencing depth planning for UMI transcriptomics
Combining RNA sequencing and pathology images identifies glioblastoma subgroups linked to survival
Stay Connected
Submit a Post to the Blog
Recent Posts
Subscribe to the RNA-Seq Blog
RNA-Seq Products & Services