

Understanding how genetic variants influence cancer is one of the major challenges in modern biomedical research. These variants can drive tumor growth, affect how a cancer behaves, and even determine how well a patient responds to treatment. In a new investigation led by researchers from Nationwide Children’s Hospital, researchers explored how looking at RNA, rather than DNA alone, can provide a deeper picture of the genetic changes that matter most in cancer.
Most genetic testing in oncology focuses on DNA sequencing, which identifies mutations in the genome. But DNA alone cannot tell us which genes the tumor is actively using. That is where RNA sequencing comes in. RNA sequencing measures the RNA molecules derived from DNA and reflects real time gene activity inside a cell. Because tumors often manipulate gene expression to their advantage, RNA based data can reveal which variants are being expressed at higher or lower levels than expected.
The research team created VarRNA, a new method that uses machine learning to classify genetic variants detected in tumor RNA. VarRNA sorts each variant into one of three categories, germline, somatic, or artifact. Germline variants are inherited, while somatic variants arise in the tumor and often contribute directly to cancer development. Artifacts are errors that appear due to technical noise. Distinguishing these categories is essential, because only true somatic variants help researchers understand tumor biology.
VarRNA data processing and variant classification pipeline
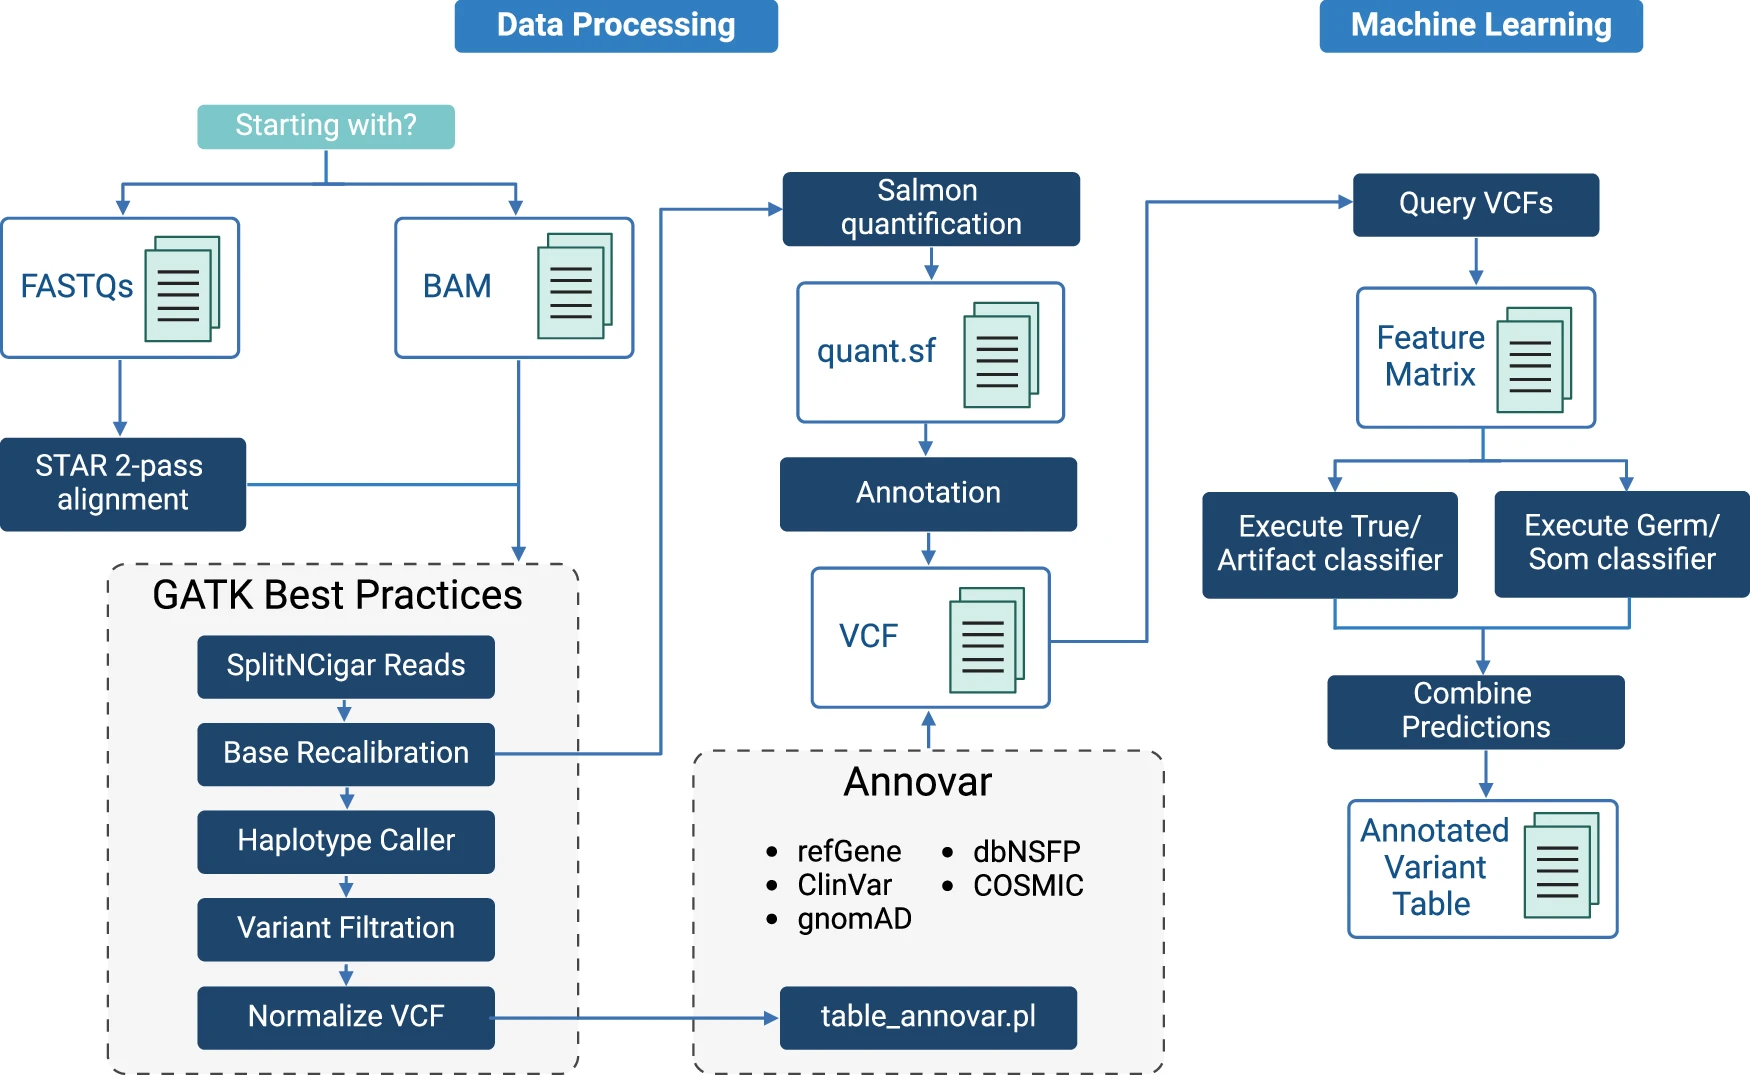
Overview of all steps performed by VarRNA using RNA-Seq data as input. The output results are an annotated variant table including classifier results for each sample. Files called out in white boxes are kept in the final output.
To develop and validate VarRNA, the team used pediatric cancer samples that had both RNA sequencing and DNA exome sequencing available. The DNA data served as a gold standard reference. By training two XGBoost models, VarRNA learned to identify patterns that indicate whether a variant is real or an artifact. This machine learning approach allowed VarRNA to outperform current RNA based variant detection tools when tested on additional cancer datasets.
One of the most striking findings is that VarRNA captured about half of the variants detected through DNA exome sequencing. Even more compelling, VarRNA uncovered unique variants present only in RNA. These RNA specific variants may reflect processes like RNA editing or tumor related expression events that are not visible in DNA. Such variants could have important roles in how cancer cells adapt and survive.
Another key insight involves allele specific expression, where one version of a gene is expressed more strongly than the other. VarRNA revealed that certain pathogenic cancer variants appear at much higher frequencies in RNA than in DNA. This imbalance can have biological consequences, especially when the variant is located in a cancer driving gene. Higher expression of a harmful variant could accelerate tumor progression or change how the cancer responds to therapies.
By highlighting differences between DNA and RNA level variant information, VarRNA offers a more complete view of tumor genetics. These results support the growing recognition that RNA sequencing can reveal clinically meaningful patterns that are not detected through DNA sequencing alone. Understanding how strongly a cancer related variant is expressed could help refine diagnoses, improve prognostic predictions, and guide treatment strategies.
VarRNA represents an important step toward integrating RNA based variant analysis into cancer research and clinical workflows. By pairing machine learning with RNA sequencing, researchers can move closer to understanding not just which variants exist in a tumor, but which ones truly matter for disease progression and patient care.
Availability – The models developed and the code used to process RNA-Seq data presented in this manuscript are open-source and available for download under the BSD 3-Clause license at https://github.com/nch-igm/VarRNA.
Bollas A, Gaither J, Schieffer KM, White P, Mardis ER. (2025) Variant calling from RNA Seq data reveals allele specific differential expression of pathogenic cancer variants. Commun Med (Lond) 5(1): 202. [article]
Related Posts





Understanding how genetic variants influence cancer is one of the major challenges in modern biomedical research. These variants can drive tumor growth, affect how a cancer behaves, and even determine how well a patient responds to treatment. In a new investigation led by researchers from Nationwide Children’s Hospital, researchers explored how looking at RNA, rather than DNA alone, can provide a deeper picture of the genetic changes that matter most in cancer.
Most genetic testing in oncology focuses on DNA sequencing, which identifies mutations in the genome. But DNA alone cannot tell us which genes the tumor is actively using. That is where RNA sequencing comes in. RNA sequencing measures the RNA molecules derived from DNA and reflects real time gene activity inside a cell. Because tumors often manipulate gene expression to their advantage, RNA based data can reveal which variants are being expressed at higher or lower levels than expected.
The research team created VarRNA, a new method that uses machine learning to classify genetic variants detected in tumor RNA. VarRNA sorts each variant into one of three categories, germline, somatic, or artifact. Germline variants are inherited, while somatic variants arise in the tumor and often contribute directly to cancer development. Artifacts are errors that appear due to technical noise. Distinguishing these categories is essential, because only true somatic variants help researchers understand tumor biology.
VarRNA data processing and variant classification pipeline
Overview of all steps performed by VarRNA using RNA-Seq data as input. The output results are an annotated variant table including classifier results for each sample. Files called out in white boxes are kept in the final output.
To develop and validate VarRNA, the team used pediatric cancer samples that had both RNA sequencing and DNA exome sequencing available. The DNA data served as a gold standard reference. By training two XGBoost models, VarRNA learned to identify patterns that indicate whether a variant is real or an artifact. This machine learning approach allowed VarRNA to outperform current RNA based variant detection tools when tested on additional cancer datasets.
One of the most striking findings is that VarRNA captured about half of the variants detected through DNA exome sequencing. Even more compelling, VarRNA uncovered unique variants present only in RNA. These RNA specific variants may reflect processes like RNA editing or tumor related expression events that are not visible in DNA. Such variants could have important roles in how cancer cells adapt and survive.
Another key insight involves allele specific expression, where one version of a gene is expressed more strongly than the other. VarRNA revealed that certain pathogenic cancer variants appear at much higher frequencies in RNA than in DNA. This imbalance can have biological consequences, especially when the variant is located in a cancer driving gene. Higher expression of a harmful variant could accelerate tumor progression or change how the cancer responds to therapies.
By highlighting differences between DNA and RNA level variant information, VarRNA offers a more complete view of tumor genetics. These results support the growing recognition that RNA sequencing can reveal clinically meaningful patterns that are not detected through DNA sequencing alone. Understanding how strongly a cancer related variant is expressed could help refine diagnoses, improve prognostic predictions, and guide treatment strategies.
VarRNA represents an important step toward integrating RNA based variant analysis into cancer research and clinical workflows. By pairing machine learning with RNA sequencing, researchers can move closer to understanding not just which variants exist in a tumor, but which ones truly matter for disease progression and patient care.
Availability – The models developed and the code used to process RNA-Seq data presented in this manuscript are open-source and available for download under the BSD 3-Clause license at https://github.com/nch-igm/VarRNA.
Bollas A, Gaither J, Schieffer KM, White P, Mardis ER. (2025) Variant calling from RNA Seq data reveals allele specific differential expression of pathogenic cancer variants. Commun Med (Lond) 5(1): 202. [article]
Related Posts
Recent Posts
 Benchmarking RNA sequencing for more accurate alternative splicing analysis
 RNA Sequencing identifies new tick-borne virus that causes flu-like illness
 Eclipsebio expands its RNA characterization capabilities with multi-attribute nanopore-based sequencing assay eSTRAND RNA QC™
 Single-cell RNA sequencing links genetic mutations to cell behavior in cancer
 Small RNA sequencing reveals regulatory roles for sdRNAs in acute myeloid leukemia
 Creative Biolabs expands advanced single-cell multi-omics and RNA sequencing solutions to support biomedical discovery
RNA-Seq Products & Services


Stay Connected